

What is ISO 10993 Certified? How to Get It ?
ISO 10993 is a series of standards for the biological evaluation of medical devices published by the International Organization for Standardization (ISO). It aims to systematically evaluate the compatibility between medical device materials and human tissues. This series of standards provides manufacturers with a globally recognized evaluation framework to identify and manage potential biological risks (such as cytotoxicity, sensitization, irritation, systemic toxicity, etc.) generated by medical devices under their intended conditions of use. A complete and compliant ISO 10993 biocompatibility test report is an indispensable key document for medical devices to complete registration declarations in major global markets.
iso 10993 test Items and Standards
The ISO 10993 series of standards consists of 20 parts, specifying concrete requirements for different risk dimensions. The selection of test items is not fixed; it is scientifically determined based on three core elements:
1.Contact Nature: Whether the device contacts the body surface, enters body cavities (blood path, tissues), or is fully implanted into the body.
2.Contact Duration: Categorized into short-term ( 24 hours), prolonged (24 hours to 30 days), and permanent (> 30 days) contact.
3.Material Composition: Different materials used in the device, such as metals, polymers, ceramics, composites, etc.
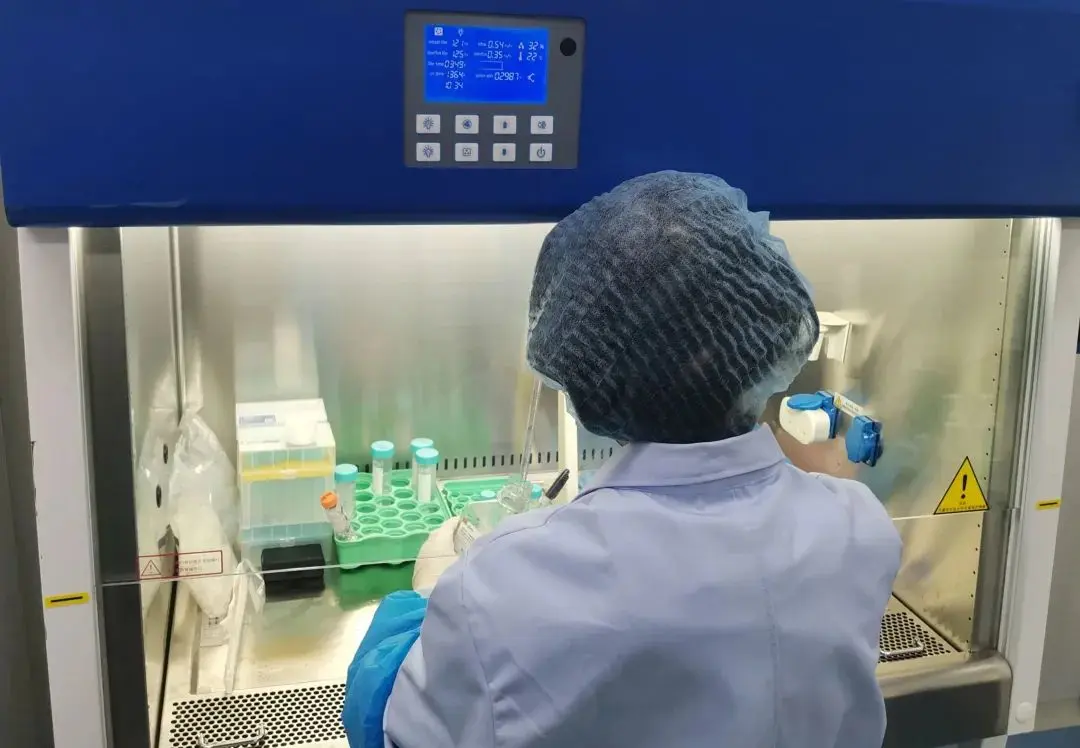
After determining the test level based on the above elements, common core test items include:
1.cytotoxicity test (iso 10993-5): Evaluates the inhibitory or destructive effects of material extracts on cell growth.
2.Skin Sensitization Test (iso 10993-10): Detects the potential of materials to cause delayed-type hypersensitivity reactions.
3.Irritation and Intracutaneous Reactivity Test (ISO 10993-10): Evaluates local irritation effects of materials on skin or tissues.
4.Acute Systemic Toxicity Test (ISO 10993-11): Evaluates adverse reactions after short-term systemic exposure to material extracts.
ISO 10993 Test Costs
ISO 10993 test costs are not fixed values; their composition is comprehensively affected by various factors:
1.Scope of Testing and Number of Items: The cost for the basic three tests (cytotoxicity, irritation, sensitization) is usually between 984.85 - 2,954.55 USD. If the product is an implantable device or contacts circulating blood, items such as genotoxicity and local reaction after implantation need to be added, driving total costs up to 15,000 USD or more.
2.Number of Samples and Variables: Different specifications, colors, material formulations, or production batches usually need to be tested as independent samples; an increase in quantity will directly push up costs.
3.Animal Testing Requirements: In vivo tests involving rats, rabbits, or larger animals (such as implantation tests) incur significantly higher breeding, operation, and ethical costs than in vitro experiments.
4.Qualifications and Reputation of the Testing Organization: Laboratories certified by CNAS and CMA, especially those complying with GLP (Good Laboratory Practice) standards, produce reports with high international recognition, and their fees are usually higher than standard commercial laboratories.
5.Service Cycle and Expedited Needs: The conventional testing cycle is 6-10 weeks. If expedited service is required (e.g., compressed to 3-4 weeks), additional rush fees will apply.
6.Standard and Regulatory Updates: It is necessary to ensure that test plans comply with the latest standard versions and target country/region-specific guidelines; otherwise, it may result in supplementary testing or duplicate work.
Optimizing Test Costs
Enterprises can effectively control test costs while ensuring compliance through proactive planning and management:
1.Early Intervention and Material Control: Introduce biocompatibility assessments during the product design stage, prioritizing mature materials with clear safe-use histories to avoid testing failures and re-testing caused by material issues later.
2.Adopting a Tiered Testing Strategy: Prioritize high-risk core items (such as cytotoxicity). If a test fails, materials can be adjusted in time to avoid ineffective investment in all items.
3.Optimizing Sample Submission Plans: Under the guidance of testing institutions, adopt the "worst-case scenario" sample principle for testing to reduce the number of unnecessary parallel samples.
4.Choosing Laboratories with International Mutual Recognition Qualifications: Although initial investments may be slightly higher, GLP reports issued by them can be directly accepted by multiple parties like the US FDA and EU Notified Bodies, avoiding duplicate testing and saving overall registration costs and time in the long run.
5.Strengthening Deep Communication with Testing Institutions: Fully provide product information before testing to jointly formulate the most economical and efficient personalized testing plan.
Scientific Management of Testing Workflow and Cycle
A standardized testing workflow typically includes the following stages:
1.Plan Formulation and Confirmation: The enterprise and laboratory jointly determine test items and standards (approx. 1-2 weeks).
2.Sample Preparation and Submission: The enterprise prepares and delivers compliant test samples (approx. 1 week).
3.Laboratory Test Execution: The core experimental phase. In vitro tests require approx. 2-4 weeks, while in vivo animal tests require 4-8 weeks or longer.
4.Data Organization and Report Preparation: The laboratory analyzes data and drafts the official report (approx. 1-2 weeks).
Enterprises are advised to incorporate biocompatibility evaluation into the overall product R&D timeline and start as early as possible. Especially for devices containing novel materials or complex designs, buffer time must be reserved for possible supplementary testing to ensure the overall market launch timeline is not affected.
Frequently Asked Questions (FAQ)
Does my medical device need to complete all test items in ISO 10993?A: No. You must select and justify the necessary combination of test items based on the risk management principles of ISO 10993-1, according to the specific contact nature, contact duration, and material of the product. This is usually determined after a risk assessment performed by the enterprise or an entrusted professional organization.
Does an ISO 10993 biocompatibility test report have an expiration date?A: The report itself does not have a fixed "expiration date." However, as long as any condition that may affect biological safety changes—such as the material source, formulation, manufacturing process, or sterilization method—a re-evaluation or supplementary testing must be conducted.
Can test reports issued by laboratories within China be used for registration in the United States or the European Union?A: It keyly depends on the laboratory's qualifications. If the laboratory is GLP-certified and its qualifications are recognized by regulatory authorities in the target market (such as FDA inspection), its reports can be accepted. To be safe, confirm the acceptability of the laboratory's qualifications with notified bodies or regulatory agencies before launching the project.
Conclusion
A deep understanding of the essence and requirements of the ISO 10993 series standards, combined with a comprehensive mastery of testing cost components, helps medical device companies formulate more precise R&D budgets, arrange more reasonable project schedules, and allocate resources more efficiently. By implementing proactive biological safety management strategies, companies can not only ensure successful regulatory reviews for their products, but also lay a solid foundation of safety for commercial success.
Email:hello@jjrlab.com
Write your message here and send it to us
What is ISO 10993 Certified? How to Get It ?
 CPSC eFiling Requirements
 Temu 2027 Children's Products and Textiles Complia
 What Compliance is Required for Exporting Adult Cl
 European Regulation PPWR Testing Services
 Squishy Toy Western Compliance Guidelines
 GB 4806.10-2025 Food Contact Coatings Testing
 Amazon Children's Jewelry DV Program Recognized TI
Leave us a message
24-hour online customer service at any time to respond, so that you worry!